 QuAcK
Ground-state energies
Build a Hartree–Fock reference and explore electron correlation with perturbation theory and coupled-cluster methods.
Ground-state methods →QuAcK
Explore and develop Green’s-function, coupled-cluster, and response methods for molecular systems.
Build a Hartree–Fock reference and explore electron correlation with perturbation theory and coupled-cluster methods.
Ground-state methods →Compute quasiparticle energies and explore charged excitations with single-shot and self-consistent GW, Green’s-function perturbation theory, and ADC.
Green’s functions →Study electronic response with the random-phase approximation and Bethe–Salpeter methods.
Response methods →Run single-shot G₀W₀ on a restricted Hartree–Fock reference with the bundled water geometry and 6-31G basis.
python docs/examples/run.py water-gw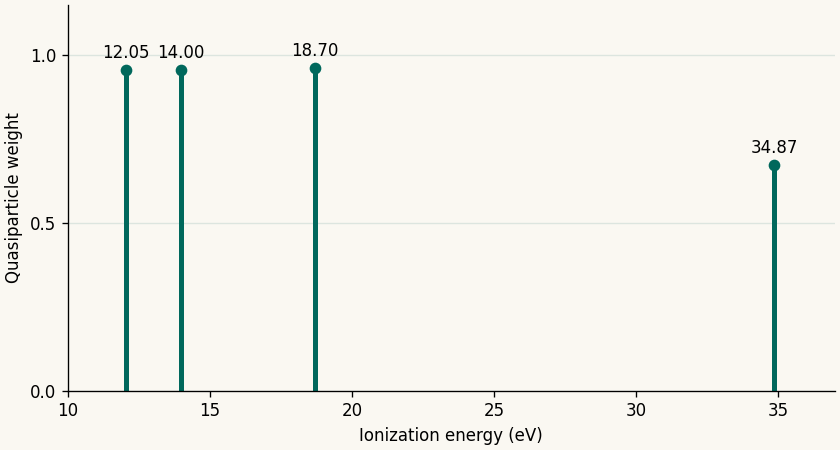
QuAcK is developed at the Laboratoire de Chimie et Physique Quantiques in Toulouse. PyDuck prepares molecular integrals with PySCF and passes them to the Fortran drivers.